GLIMPSE2 SV Imputation Overview
| Pipeline Version | Date Updated | Documentation Author | Questions or Feedback |
|---|
| See changelog for version information. | See changelog | Terra Scientific Pipeline Services | Please file an issue in WARP. |
Introduction to the GLIMPSE2 SV Imputation pipeline
The GLIMPSE2 SV Imputation pipeline imputes structural variants, SNPs, and INDELs from a manifest of input GVCF paths (or from GVCF/index arrays that are converted to a manifest). It uses GLIMPSE2-based phasing, panel-informed bubble processing, and cohort-aware merge/re-annotation to produce final per-contig imputed VCF outputs.
GLIMPSE2 SV Imputation Summary
The Glimpse2SVImputation workflow is a WDL-based pipeline for structural variant imputation using GLIMPSE2.
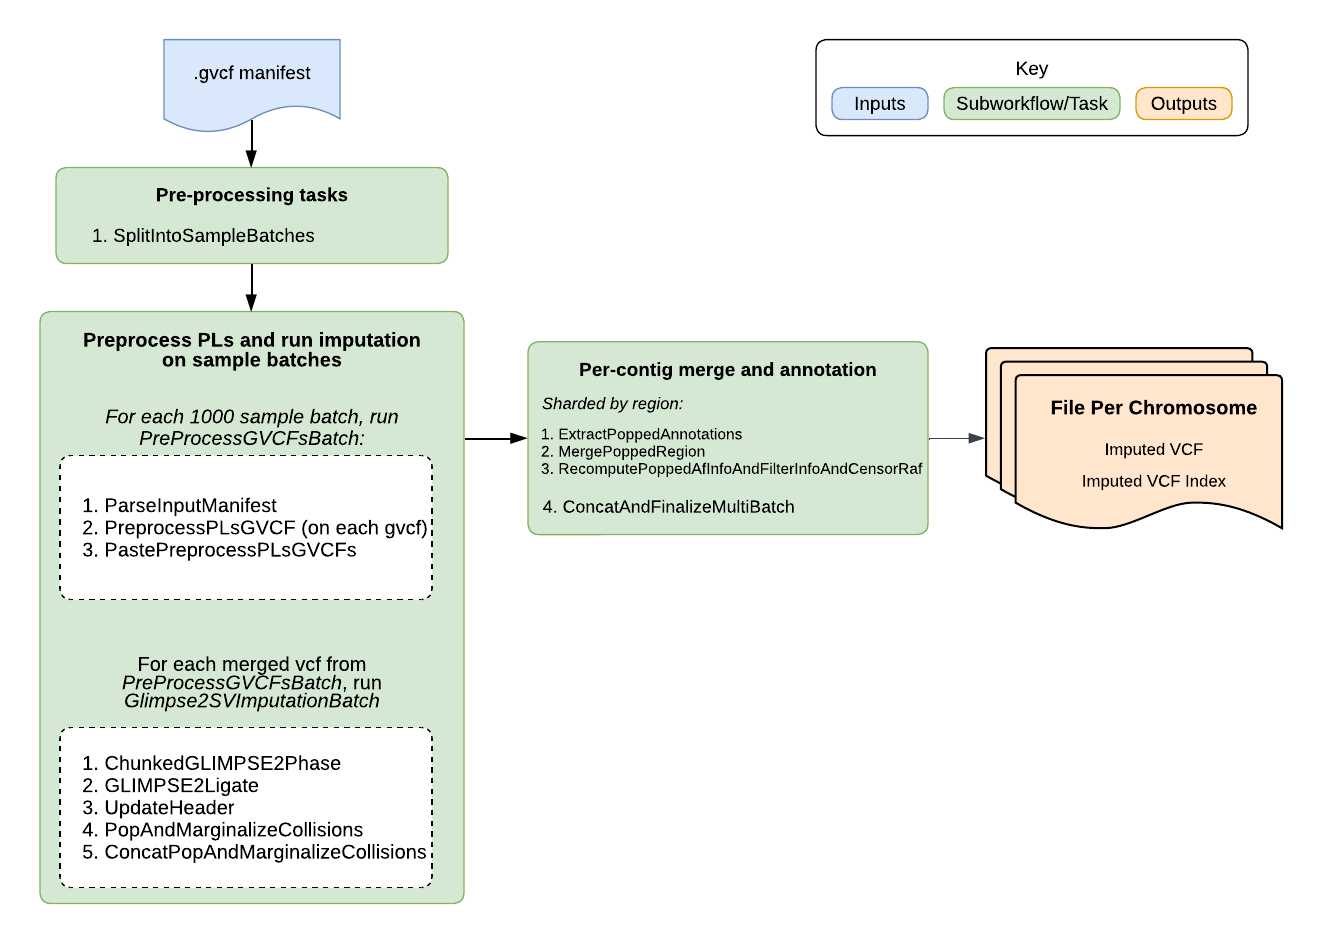
This top-level workflow acts as a gateway that scales to large cohorts by splitting samples into batches, running preprocess + batch imputation subworkflows per batch, then merging per-batch results back into cohort-level per-contig VCFs.
The workflow processes requested chromosomes independently, extracts/reformats bubble likelihoods, phases and ligates imputed chunks, pops/marginalizes collisions, merges batch sample columns, recomputes AF/INFO across all samples, optionally filters by INFO threshold, optionally censors values of RAF below threshold, and indexes final outputs.
Pipeline Features
| Pipeline features | Description | Source |
|---|
| Assay type | Structural variant imputation using GLIMPSE2 | GLIMPSE2 |
| Overall workflow | Manifest normalization, preprocessing, chunked phase/ligate, pop/marginalize, merge, and AF/INFO re-annotation | Defined in Glimpse2SVImputation.wdl + imported subworkflows/tasks |
| Workflow language | WDL 1.0 | openWDL |
| Sub-workflows | Gateway workflow + PreprocessPLsGVCF + Glimpse2SVImputationBatch | Imported from sibling WDLs in sv_imputation/ |
| Genomic processing | Contig-by-contig processing with nested shard/region scatters | Workflow scatter logic |
| Cohort scalability | Input manifest splitting via sample_batch_size, then batch-level contig merge | Gateway orchestration |
| Algorithms | GLIMPSE2 phase/ligate + custom paste/concat + cohort AF/INFO recomputation | Task commands in batch/task WDLs |
| Data input file format | GVCF/GVCF index manifest or GVCF/index arrays | Workflow input block |
| Data output file format | Per-contig imputed VCFs with index files | Workflow outputs |
| Containers | GLIMPSE2, bcftools/samtools, Python, custom SV tooling containers | Runtime blocks |
| Resource optimization | Parallelization by sample batch, chromosome, chunk, and pop region | Workflow architecture |
This gateway workflow expects manifest- or array-based GVCF inputs plus SV panel/chunk resources.
| Input | Description |
|---|
input_gvcfs | Optional array of input GVCF paths. If both arrays and a manifest are supplied, arrays take precedence. |
input_gvcf_idxs | Optional array of matching GVCF index paths for input_gvcfs. |
gvcf_manifest | Optional manifest TSV used when input arrays are not provided. The workflow expects the manifest to contain gvcf_path and gvcf_index_path columns. |
sample_batch_size | Number of samples per batch for gateway-level scaling (default: 1000). |
output_basename | Basename used for intermediate and final outputs. |
preprocess_panel_bubble_split_sites_only_vcf | Panel VCF used in the preprocessing stage for bubble extraction. |
preprocess_panel_bubble_split_sites_only_vcf_idx | Index for preprocess_panel_bubble_split_sites_only_vcf. |
extract_bubble_likelihoods_extra_args | Optional extra arguments passed to the preprocessing step. |
paste_regions | Regions passed to the hierarchical merge in the preprocessing step. |
chromosomes | Chromosomes/contigs to process. |
genetic_maps_tsv | TSV mapping contig names to GLIMPSE2 genetic map files. |
ref_dict | Reference dictionary used in header updates. |
chunked_panel_json | JSON describing chunk bins and regions by chromosome. |
extra_phase_args | Command-line arguments passed to GLIMPSE2_phase. Default value is the default in the workflow input block. |
glimpse_phase_cpu_override | Optional CPU override for GLIMPSE2 chunk phasing. If unset, the batch workflow defaults to 4 threads. |
pop_glimpse2_panel_resources_json | JSON containing panel resources used in pop/marginalize processing. |
info_filter_for_inclusion | Optional INFO threshold for variant inclusion; default 0.0 (disabled). |
raf_censoring_threshold | Remove RAF annotations for sites with RAF <= this value; default 0.0 (disabled). |
pipeline_header_line | Optional additional header line added to output VCF headers. |
glimpse2_docker | Docker image used for GLIMPSE2 phases and ligation tasks. |
Workflow Tasks
The top-level workflow orchestrates batching, per-batch preprocessing/imputation, and contig-level merge/finalization.
| Task / Call | Purpose | Input Dependencies | Key Function |
|---|
ConvertInputArraysToManifest | Convert input GVCF/index arrays into a manifest | input_gvcfs, input_gvcf_idxs | Supports large submissions via manifest abstraction |
SplitIntoSampleBatches | Split the manifest into sample-level batches | gvcf_manifest_to_use, sample_batch_size | Enables gateway-level cohort scaling |
PreProcessGVCFsBatch (PreprocessPLsGVCF) | Preprocess one batch of GVCFs into a merged BCF | Batch manifest + preprocessing resources | Produces one preprocessed BCF/index per batch |
RunBatch (Glimpse2SVImputationBatch) | Run per-batch chunked SV phase/ligate/pop processing | Preprocessed BCF + chromosome/map/chunk/panel resources | Produces per-batch per-contig popped posterior BCFs |
ExtractPoppedAnnotations | Extract AF/INFO annotations from each batch contig BCF in shard regions | Per-batch-and-contig popped BCF + index | Captures per-batch-and-shard annotations for cohort recomputation |
MergePoppedRegion | Merge sample columns across batches for each shard | Arrays of per-batch-and-shard VCFs | Creates the full-cohort shard VCF |
RecomputePoppedAfInfoAndFilterInfoAndCensorRaf | Recompute AF/INFO across the merged cohort and re-annotate for each shard and optionally filter by INFO and censor RAF | Merged per-shard VCFs + annotations + sample counts + INFO and RAF thresholds | Restores cohort-correct AF/INFO values |
ConcatAndFinalizeMultiBatch | Concat shards and finalize VCF/index outputs | Per-shard re-annotated+filtered+censored VCFs | Emits imputed_vcf and imputed_vcf_index per contig |
FilterInfoAndCensorRafAndFinalizeBcf | Optionally filter by INFO and censor RAF and then finalize VCF/index outputs for single batches | Popped contig BCF | Emits imputed_vcf and imputed_vcf_index per contig |
Outputs
Upon successful completion, the workflow emits final per-contig imputed outputs and matching indexes.
| Output | Description |
|---|
imputed_vcfs | Final per-contig imputed VCF files |
imputed_vcf_indexes | Index files for final per-contig imputed VCF files |
PreprocessPLsGVCF summary
The PreprocessPLsGVCF workflow preprocesses each input GVCF in a batch and hierarchically merges all preprocessed outputs into a single batch-level BCF for downstream GLIMPSE2 phasing.
PreprocessPLsGVCF internal processing
| Step | Purpose |
|---|
ParseInputManifest (ParseVcfManifestIntoArrays) | Parse the batch manifest into arrays of GVCF paths and index paths |
PreprocessPLsGVCF scatter (PreprocessPLs) | Run per-sample extract-bubble-PLs to generate preprocessed BCFs |
PastePreprocessPLsGVCFs (MultilevelHierarchicallyMergeVcfs) | Hierarchically merge preprocessed BCFs across configured regions |
PreprocessPLsGVCF outputs
| Output | Description |
|---|
preprocessed_pls_bcf | Batch-level merged preprocessed BCF |
preprocessed_pls_bcf_idx | Index for preprocessed_pls_bcf |
num_samples | Number of samples represented in the batch |
Glimpse2SVImputationBatch summary
The Glimpse2SVImputationBatch workflow is the per-batch subworkflow used by Glimpse2SVImputation. It takes one preprocessed batch BCF and runs chromosome-level chunked phasing, ligation, header normalization, and pop/marginalize post-processing.
Batch internal processing
| Step | Purpose |
|---|
ChunkedGLIMPSE2Phase | Run GLIMPSE2_phase over each chunk for each chromosome |
GLIMPSE2Ligate | Ligate chunk-level phased outputs into one chromosome-level BCF |
UpdateHeader | Rebuild/update headers with dictionary and optional pipeline line |
PopAndMarginalizeCollisions | Apply pop/marginalize processing over one or more regions per chromosome |
ConcatPopAndMarginalizeCollisions (ConcatBcfs) | Concatenate regional popped BCFs into one popped BCF per chromosome |
Batch outputs
| Output | Description |
|---|
glimpse2_bubble_posteriors_vcf | Per-chromosome post-ligation BCFs with updated headers |
glimpse2_bubble_posteriors_vcf_idx | Indexes for bubble posterior outputs |
glimpse2_popped_posteriors_vcf | Final per-chromosome popped BCFs for upstream cohort merge |
glimpse2_popped_posteriors_vcf_idx | Indexes for popped posterior outputs |
MultilevelHierarchicallyPasteVcfsStreaming summary
The MultilevelHierarchicallyMergeVcfs workflow provides scalable multi-level merging for squared-off single-sample VCF/BCF inputs. It performs region-first hierarchical batching, optional streaming/localization, and final region concatenation.
MultilevelHierarchicallyMergeVcfs internal processing
| Step | Purpose |
|---|
CreateBatches (L0/L1/L2) | Split input files into hierarchical batch groups |
MergeVcfs (L0/L1/L2) | Merge each batch within each region with optional streaming timeouts |
FinalRegionMerge | Collapse remaining per-region intermediate files when needed |
ConcatBcfs | Concatenate finalized region outputs into one merged BCF |
MultilevelHierarchicallyMergeVcfs outputs
| Output | Description |
|---|
merged_bcf | Final merged BCF across all input samples and regions |
merged_bcf_idx | Index for merged_bcf |
Glimpse2SVImputationQuotaConsumed summary
The QuotaConsumed workflow computes submitted sample count for service quota accounting from gvcf_manifest.
QuotaConsumed internal processing
| Step | Purpose |
|---|
CountGvcfsFromManifest | Count GVCF entries in the manifest and report quota_consumed |
QuotaConsumed outputs
| Output | Description |
|---|
quota_consumed | Number of submitted GVCF entries counted from the manifest |
Glimpse2SVImputationQC summary
The InputQC workflow validates manifest-level and GVCF-level readiness before SV imputation.
- required manifest columns are present:
gvcf_path, gvcf_index_path - manifest has at least one row and no empty required values
- GVCF paths are unique
- GVCF and index extensions are valid (
.vcf.gz/.gvcf.gz and .tbi) - GVCF and index basenames are matched
- paths use valid
gs:// format and are accessible - GVCF file size is below the configured threshold (default
10 GB) - GVCF headers are compatible with the expected reference dictionary
- each GVCF contains exactly one sample
- sample IDs are unique across all provided GVCFs
- PL and GT FORMAT IDs are present in each GVCF header
- each GVCF is vcf version 4.x
| Output | Description |
|---|
passes_qc | Overall QC pass/fail status |
qc_messages | Aggregated QC failure messages (empty when QC passes) |
Important notes
- Runtime parameters are optimized for Broad's Google Cloud Platform implementation.
Help us make our tools better by filing an issue in WARP; we welcome pipeline-related suggestions or questions.